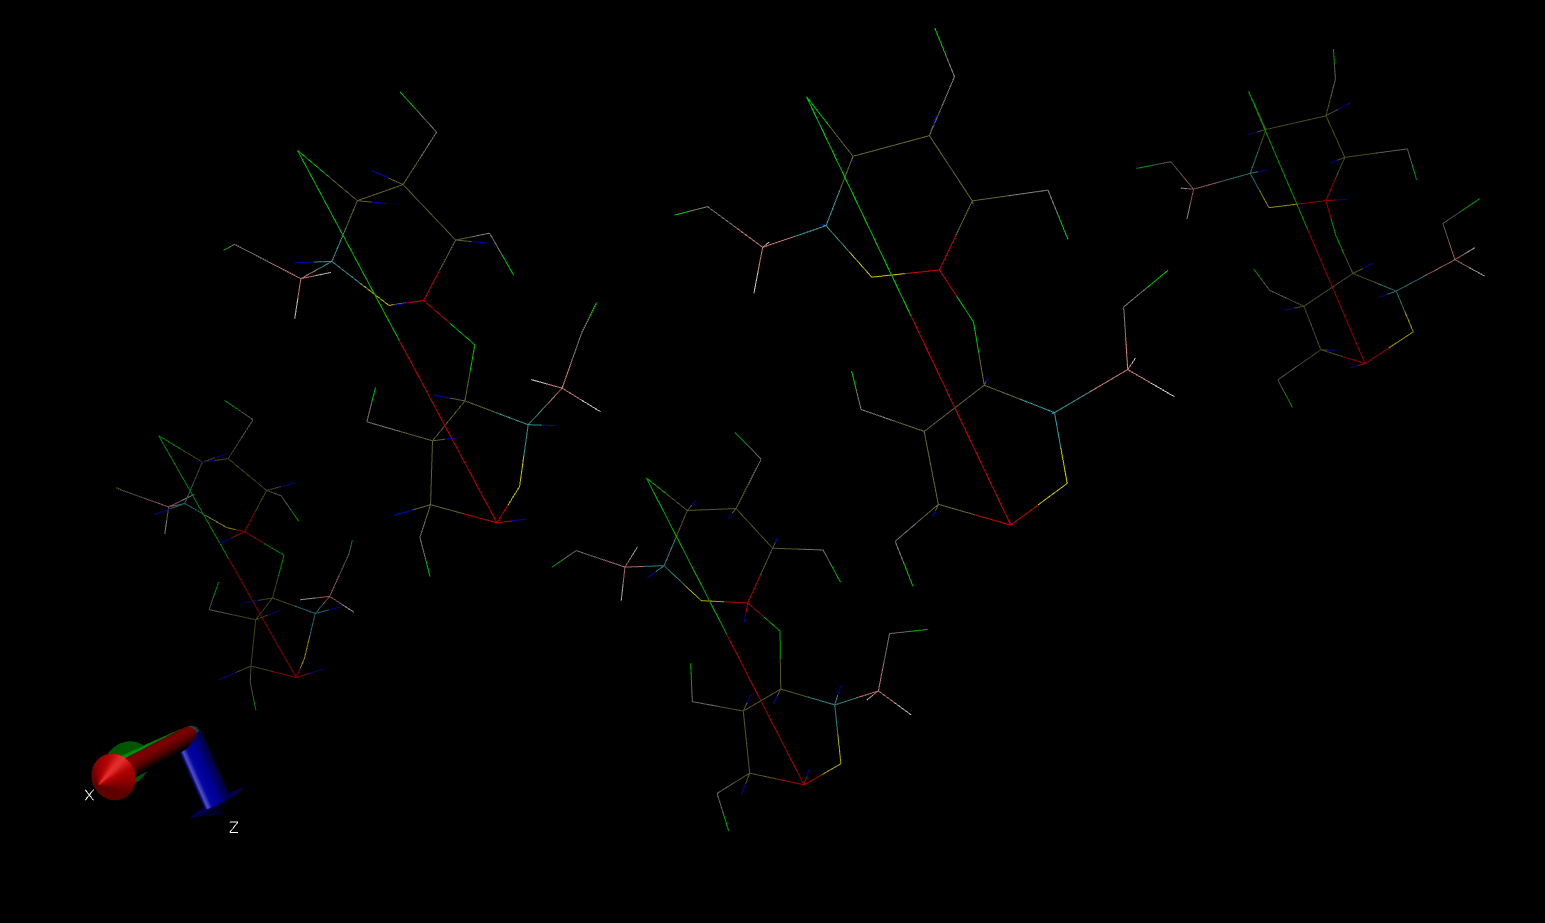
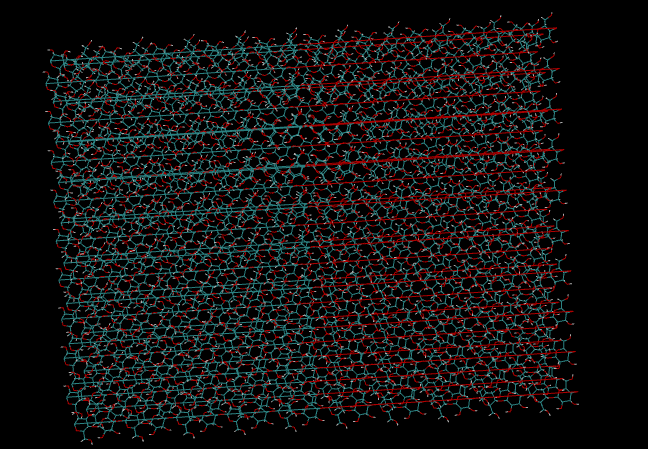
As part of the conversion process, the files top-all36-carb.rtp and top-all36-carb.prm are required. I attempted to use versions available online, specifically from this GitHub repository:
However, when I use these files, an extra bond appears within the cellulose structure, which is clearly incorrect.
Could someone kindly share the correct or working versions of top-all36-carb.rtp and top-all36-carb.prm that were used successfully for this cellulose system? Or advise on how to modify the existing ones to avoid the incorrect bond generation?
That is not a tutorial, but just another user like you asking for help
I am not so sure about that. You have a periodic system and the long bond like this would show up in VMD this way for bonds reaching across periodic boundaries. It appears, however, that your box dimensions are not proper for that and there are significant gaps between periodic images.
I don’t think those are the problem here.
Those are for certain not the original files. The canonical location for obtaining the official CHARMM force field parameters is on Alex Mackerell’s homepage.
Warning: bond parameter 1 for [NC311 NC311 ] was not found
Info: converting angles
Warning: angle parameter 1 for [NC311 NC311 NC311 ] was not found
Info: converting dihedrals
Warning: dihedral parameter 1 for [NC311 NC311 NC311 NC311 ] was not found
Info: converting impropers
Info: conversion complete
this is the output when iam using updated version of .prm and .rtf files
##########################################################################
 Charmm36_FF/toppar at master · ChayaSt/Charmm36_FF · GitHub